Methane, CH₄, has four identical bonds pointing towards the corners of a tetrahedron, 109.5° apart, as the molecular geometry page shows. However, on the electron cloud page, the orbitals available to carbon’s outer electrons are one round s orbital and three p orbitals at right angles to one another. None of these points towards the corners of a tetrahedron. So how can four identical bonds come from orbitals that are not identical? Hybridisation answers this by combining the original orbitals into new ones.
“Combining orbitals” may sound like piling two clouds together, but the colours carry another piece of information: the values being combined have positive and negative signs. They are wave-function values, not positive and negative electric charges on the electron. Pick the same position in both descriptions and see how the values add: matching signs reinforce, while opposite signs reduce or cancel one another. One side of a shape growing and the other shrinking now has a cause we can follow. We can then include more orbitals and compare sp, sp² and sp³ directions, returning throughout to the bonds that prompted the model.
01 · Add a wave, not an electron
We need a description of where an electron may be found. An orbital gives a wave-function value ψ at each position, a number that varies from place to place and carries a sign; the probability rule of quantum theory makes the local probability density ∣ψ∣2. In a small volume, density multiplied by that volume gives the approximate probability. This rule is a starting point of our quantum model, not a claim that an electron follows a visible path. The real functions used here can be positive or negative, but their squares cannot.
The s map has the same amplitude at equal distances in every direction, so its cloud is spherical. A px map changes sign across the nucleus and is zero on the plane between its two lobes. Both lobes belong to one function. To describe a bonding direction, we want one side to become larger than the other. Adding s to px can do this: one side reinforces while the other cancels. The two numerical examples below show how the addition changes the values. Applying the same operation at every position builds the full three-dimensional function.
What exactly are we adding?
A = the signed amplitude contributed by the s orbital at one chosen position. B = the signed amplitude contributed by the pₓ orbital at that same position. These are two numbers, not two atoms or two electrons. Here we choose positions where their magnitudes match and display them in relative units.
Positive side of pₓ
A: s contribution = +1 B: pₓ contribution = +1
Adding the amplitudes gives +1 + (+1) = +2. The density is the square of this combined value: 2² = 4. Adding the original densities would give 1² + 1² = 2, which misses this interference. These relative values still need the overall normalisation explained next.
Negative side of pₓ
A: s contribution = +1 B: pₓ contribution = −1
Adding the amplitudes gives +1 + (−1) = 0. The density is the square of this combined value: 0² = 0. So the density at this position is zero. The values of a function have cancelled; two electrons have not been destroyed.
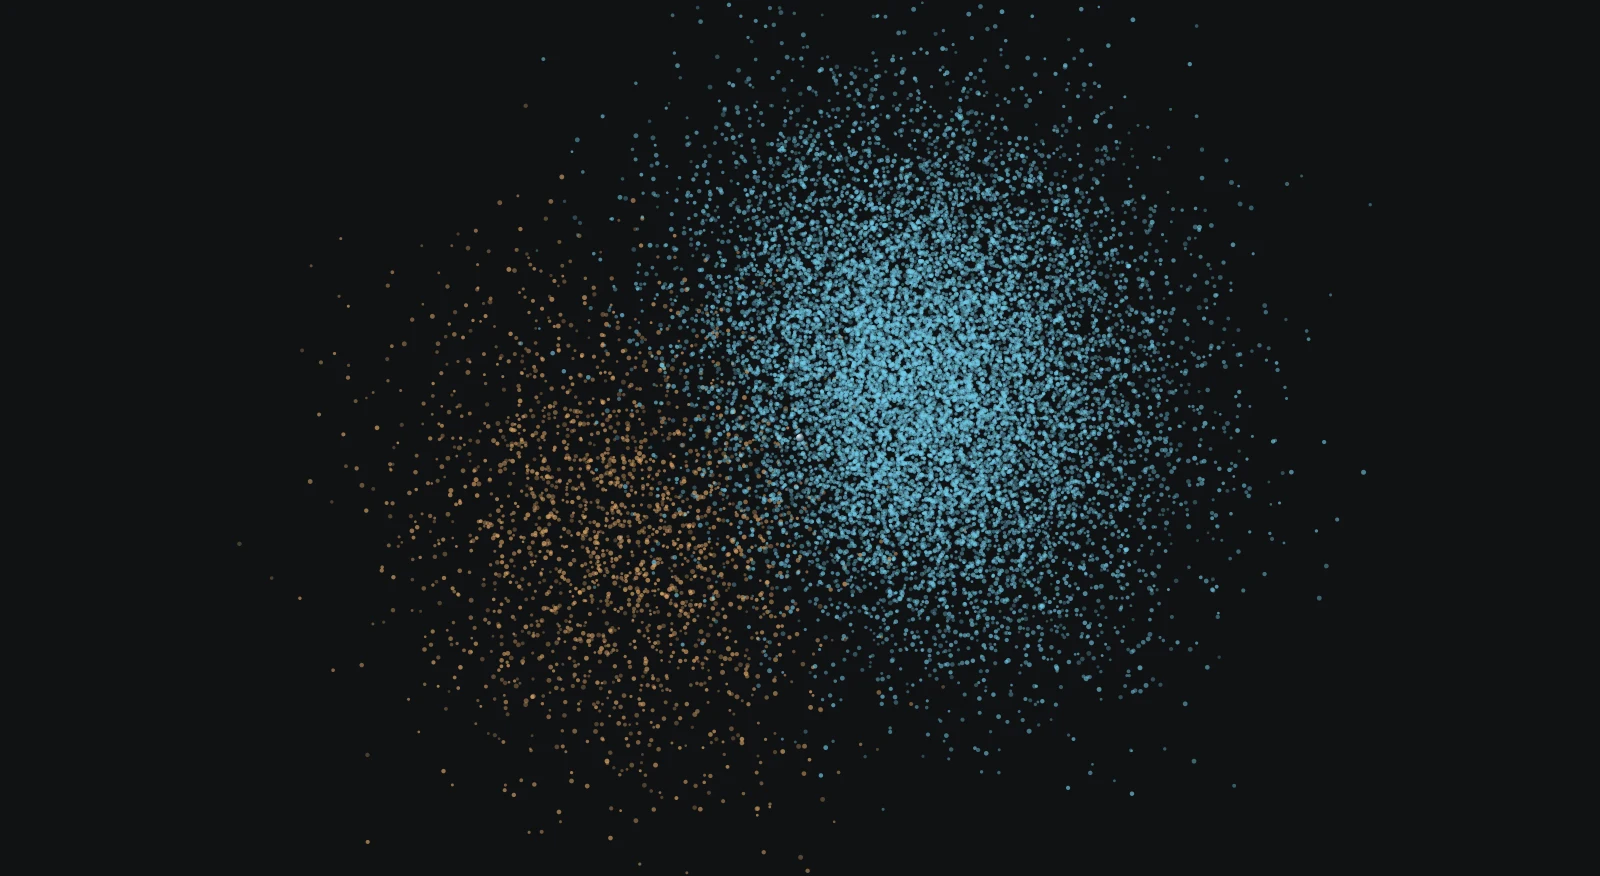
Blue marks positive phase and amber marks negative phase; neither colour represents electric charge. Adding two negative values can reinforce the amplitude too. Subtracting an orbital means adding its opposite, so both lobes change sign. The tiles show signed values, not particles being destroyed.
02 · Building sp, sp2 and sp3 one orbital at a time
We can use the same method to build a family of hybrid orbitals. Choose sp, sp2 or sp3 and press “Build step by step”. The two orbital maps move onto the same nucleus, because we add amplitudes belonging to the same positions in space. At each position, the values combine. Where the maps have the same phase, the amplitude becomes larger in magnitude; where the phases are opposite, cancellation reduces it. The displayed cloud changes with these values. For sp3, the construction combines s with px and keeps that result visible while py and pz are added one at a time. This lets us inspect what each contribution changes. The full combination could also be written as one expression; the order shown is a teaching choice. The x, y and z axes pass through the nucleus and help us identify directions after rotating the view. Each h button builds another member of the family, a hybrid orbital pointing in a different direction.
2 original orbitals → 2 hybrid orbitals. We build one member at a time and keep the running result between additions. The member selected here is h1, where h is simply the name we give a hybrid orbital.
h1=0.707s+0.707px
The coefficients set the weight and the sign of each contribution. A minus sign flips the phase of a whole orbital, which does not delete any of its electrons. Each intermediate shape is normalized, so that the total probability stays equal to 1.
A Already built · sB Joining now · +pₓ
Positive phase Negative phaseGold dot: one nucleus · gold ring: sampled position · drag to orbit
First inspect A on the left: the spherical s orbital. B on the right is the pₓ contribution. We show just these two maps.
What is changing at the gold ring?
A is the amplitude of the retained s contribution at this exact position. B is the weighted pₓ contribution at the SAME position, including its sign. The animation turns B on from 0% to 100%; only after adding do we square to obtain density.
A · s+0.402
B · pₓ × 0%0.000
Amplitude: A + B+0.402
Density: (A + B)²0.1615
These are signed values at a position, not electron counts. A probe is a mathematical readout, not a quantum measurement. The apparent movement of the maps is a teaching arrangement, not an actual time evolution of an atom.
Return to the numbers used in this construction
We have seen how signs make two contributions reinforce or cancel. Now we need to choose their sizes. Otherwise adding two complete orbital maps would change the total probability. The numbers multiplying s and p are called coefficients; they scale the amplitude, not the electron count.
Our starting s, pₓ, pᵧ, and p𝓏 functions are normalised: the sum of each squared amplitude over all space is 1. We write that sum as an integral, ∫. They are also orthogonal, meaning that the integral of the product of two different functions is zero. For example, s has the same sign at x and −x while pₓ changes sign, so the two equal and opposite products cancel when all positions are included. The same reflection argument works for each pair of different p directions.
Let h = cₛs + cₓpₓ + cᵧpᵧ + c𝓏p𝓏. Squaring produces the four squared terms and products between different functions. Every mixed product sums to zero, while each original squared function sums to 1. This leaves the squared coefficients. To keep total probability 1, we therefore require:
∫h2dV=cs2+cx2+cy2+cz2=1
We also want different members of the family to remain orthogonal, so that we have separate orbital descriptions. Multiplying hᵢ by hⱼ and applying the same cancellation gives a second rule: multiply corresponding coefficients and add; the result must be zero. These two rules will determine the numbers below.
For sp we want two equivalent directions along the x axis. One points right and one left, so only the sign of the pₓ contribution changes. Write h₁ = cs + bpₓ and h₂ = cs − bpₓ, with c and b positive. The probability rule gives c² + b² = 1; the orthogonality rule gives c² − b² = 0. Adding the equations eliminates b².
2c2=1⟹c=21,b2=1−21=21⟹b=21
h1=2s+px,h2=2s−px,21≈0.707
The s contribution is c² = 1/2 = 50% of the normalisation. The two directions are opposite along one line, so their angle is half a full turn: 360°/2 = 180°. Only one of the three p directions was used, leaving 3 − 1 = 2 unchanged.
During construction, only part of an incoming function has been added. If its current coefficients are wₛ, wₓ, wᵧ, w𝓏, the same squared-sum argument gives total probability D = wₛ² + wₓ² + wᵧ² + w𝓏². Divide the whole amplitude by √D: squaring then divides probability by D, giving D/D = 1. For example, the unfinished combination s + 0.5pₓ has D = 1² + 0.5² = 1.25, so we display (s + 0.5pₓ)/√1.25. This keeps intermediate shapes comparable.
sp · 180°
1 s + 1 p → 2 hybrid orbitals, pointing in opposite directions. Two p orbitals remain unhybridized. In the usual model of ethyne, they allow two perpendicular π bonds.
sp² · 120°
1 s + 2 p → 3 coplanar hybrid orbitals. One p orbital remains perpendicular to the plane. In ethene, side-on overlap of those p orbitals supplies the π bond.
sp³ · 109.47°
1 s + 3 p → 4 tetrahedral hybrid orbitals. Methane uses four equivalent bonding directions. Real bond angles can depart from this ideal when lone pairs or different substituents are present.
Model scope: normalized Gaussian s/p functions illustrate angular shape and interference; atomic radial nodes are omitted. Point-cloud samples follow ∣ψ∣2; neither a dot nor the edge of the visible cloud is a literal electron or a hard boundary. Hybridization is a bonding model, not an ab initio prediction of molecular geometry.
Check your understanding: does sp3 create four new electrons?
No. Four original orbitals are replaced by four linear combinations. Electron number is unchanged; occupation depends on the atom and its bonding environment.